Observables
An observable is a quantity computed from every simulation of a run: a timecourse, such as a concentration in ng/ml or normalized to its peak, or one value per simulation, such as the peak itself or the area under the curve. You pass them to Simulator.run(model, scan, observables). There are three types:
Formula: an expression of model variables, with reductions over time such asmaxandmean.PK: the non-compartmental analysis of a concentration with pkpdutils, e.g.cmax,aucand the half-life.Custom: your own python function.
Observables are evaluated on the full solution of every simulation, before the result is interpolated onto a time grid, so they are as exact as the simulation itself.
Formulas and reductions
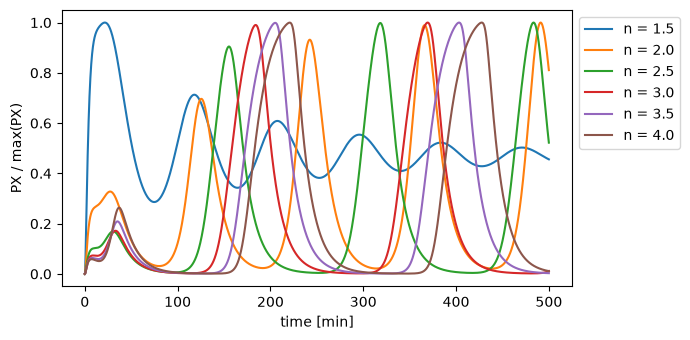
A Formula uses the math of PEtab. It reads the variables of the model (S is the amount of a species, [S] its concentration, plus time, parameters and reactions) and other observables by their id. Here the repressilator runs for six values of the Hill coefficient n, a scan with one dimension, see Parameter scans:
import matplotlib.pyplot as plt
import numpy as np

from sbmlsim.plot import label_axes
from sbmlsim.resources import REPRESSILATOR_SBML
from sbmlsim.simulation import Dimension, Formula, Scan, Simulation
from sbmlsim.simulator import Simulator

simulator = Simulator()
scan = Scan(
    Simulation(end=500),
    [Dimension("hill", values={"n": np.linspace(1.5, 4.0, 6)})],
)
observables = [
    Formula("px", "PX"),
    Formula("px_max", "max(px)"),
    Formula("px_mean", "mean(px)"),
    Formula("px_250", "at(px, 250)"),
    Formula("px_rel", "px / px_max"),
]
res = simulator.run(REPRESSILATOR_SBML, scan, observables)
print(res.q["px_max"].round(0))
print(res["px_rel"].dims)
px_max has one value per point of the scan, the number of PX molecules at the peak. px_rel divides a timecourse by a value per simulation, so every simulation is normalized to its own maximum. Every simulation has its own time points, the steps of the integrator, so px_rel is over _point and res.sel(hill=k) selects the timecourse of one point:
fig, ax = plt.subplots(figsize=(7, 3.5))
for k, n in enumerate(res["n"].values):
    point = res.sel(hill=k)
    ax.plot(point.q["time"], point.q["px_rel"], label=f"n = {n:.1f}")
label_axes(ax, "time", "PX / max(PX)")
ax.legend(loc="upper left", bbox_to_anchor=(1, 1))
fig.tight_layout()
A reduction turns the timecourse of each simulation into one value:
| reduction | value |
|---|---|
max(x), min(x) |
the largest and the smallest value, ignoring NaN |
mean(x) |
the time-weighted mean: the integral (trapezoidal rule) divided by the simulated time span |
at(x, t) |
the value at time t, interpolated linearly; NaN outside of the simulated times |
A formula that reads only values per simulation is itself a value per simulation, otherwise it is a timecourse. A value per simulation inside a timecourse is the same at every time point, which is how px / px_max normalizes to the maximum and ins / at(ins, 0) to the baseline. With two or more arguments, max and min are the elementwise maximum and minimum of PEtab.
The t of at is a number in the time unit of the model, or a value per simulation with that unit, such as pk.tmax. At the time of a change or an event, at gives the value after it. Neither the padding of a ragged scan nor the steady state after the end count as time points of a reduction.
Units
The unit of a formula is derived from the units of what it reads, and a reduction keeps the unit of its argument. With unit= the formula is converted into that unit, and the observable has this unit everywhere: in the result and in every observable that reads it.
from sbmlsim import Q
from sbmlsim.resources import MIDAZOLAM_SBML
from sbmlsim.simulation import PK, Change, Custom

simulation = Simulation(
    time_unit="hr",
    end=24,
    changes=[Change(0, {"PODOSE_mid": Q(7.5, "mg")})],
)
mass = Formula("mid", "[Cve_mid] * Mr_mid", unit="ng/ml")
res = simulator.run(MIDAZOLAM_SBML, simulation, [mass, Formula("mid_max", "max(mid)")])
print(res.units["mid"], res.units["mid_max"])
[Cve_mid] is the molar concentration of midazolam in the venous plasma and Mr_mid its molar mass, so mid is the mass concentration in ng/ml.
Some rules for units:
- A formula that mixes units of one dimension at different scales raises when the scan is compiled, e.g.
mid - [Cve_mid] * Mr_midwithmidin ng/ml and the product in mg/l. Declare one of them as aFormulawithunit=; a factor in the formula does not convert it. - pint cannot derive a unit for a comparison or a
piecewise. Such a formula needsunit=, which is taken as given:Formula("high", "piecewise(1, mid > 100, 0)", unit="dimensionless"). Mixed scales inside it are not checked, sopiecewise(mid, time < 600, [Cve_mid] * Mr_mid)runs and is off by a factor of 1000 in one branch. - A formula without a unit it needs, a unit that cannot be converted, an unknown symbol and observables that read each other in a cycle all raise before any simulation runs.
PK
PK runs a non-compartmental analysis of a timecourse with pkpdutils. Every PK parameter is one value per simulation, named <id>.<parameter>. The analysis uses the unit of the timecourse, so with mid in ng/ml the concentrations are in ng/ml and the times in minutes, the time unit of the model:
doses = Scan(
    simulation,
    [
        Dimension(
            "dose",
            values={"PODOSE_mid": Q([5.0, 7.5, 15.0], "mg")},
            labels=["5 mg", "7.5 mg", "15 mg"],
        )
    ],
)
observables = [
    mass,
    PK("pk", "mid", dose="PODOSE_mid", route="oral"),
    Formula("auc_per_cmax", "pk.auc_inf_obs / pk.cmax"),
]
res = simulator.run(MIDAZOLAM_SBML, doses, observables)
print(res.q["pk.cmax"].round(2))
print(res.q["auc_per_cmax"].to("hr").round(2))
A formula reads a PK parameter by its name: auc_per_cmax divides two of them and barely changes with the dose. res.q[...] gives a variable with its unit, which .to(...) converts.
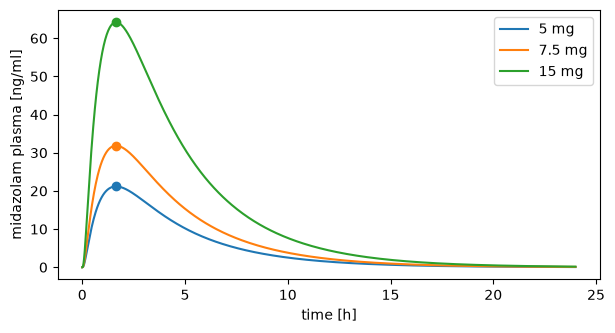
The peak of every curve is at (pk.tmax, pk.cmax):
fig, ax = plt.subplots(figsize=(7, 3.5))
for label in res["dose"].values:
    point = res.sel(dose=label)
    (line,) = ax.plot(point.q["time"].to("hr"), point.q["mid"], label=label)
    ax.plot(point.q["pk.tmax"], point.q["pk.cmax"], "o", color=line.get_color())
label_axes(ax, "time", "midazolam plasma")
ax.legend()
res.nca("pk") returns the NCAResult of pkpdutils and res.to_timecourses("mid") its Timecourses, for further analysis with pkpdutils.
How the doses are found:
- The values the simulation assigns to the dose target are the doses, and their times are the dose times. A dose of a scan dimension and multiple dosing with
Change([0, 12], ...)therefore need nothing more. A value set before the start is a dose at the start. - A value of zero is no dose. A control point with a zero dose gets
NaNfor the parameters that need a dose (e.g.pk.cl_f), and the other points keep them. dose=may also be a quantity, a fixed dose at the start. Without a dose, the parameters that need one (clearance, volume) are left out.
The parameters are the ones pkpdutils derives for the route and dosing, e.g. pk.cmax, pk.tmax, pk.auc_inf_obs, pk.thalf, pk.cl_f and pk.flags, the flags of the analysis. parameters=[...] keeps a subset and options=pkpdutils.NCAOptions(...) changes the settings of the analysis. At a change after the start the timecourse has two rows, the states before and after it; the analysis sees the state after it. The analysis uses the time points of the simulation. The steps of the integrator follow the curve within the tolerances of the integrator, so they need no time grid; times of the simulation gives the analysis the times of a study instead, e.g. its sampling times.
Custom functions
A Custom observable calls a function function(time, values) once per simulation. time holds the time points in the time unit of the model and values a dict with the timecourses of the symbols, each in its unit. A time appears twice at a change after the start or an event (the states before and after it). The function returns a float, or an array of the length of time with kind=ObservableKind.TIMECOURSE (ObservableKind is in sbmlsim.simulation). You give the unit of the result.
The function must be defined at the top level of a module, so that it can be sent to the worker processes of a scan; a lambda or a closure raises. examples/observables.py has time_above, the time during which mid (in ng/ml) is above 25 ng/ml:
from examples.observables import time_above

res = simulator.run(
    MIDAZOLAM_SBML,
    doses,
    [mass, Custom("t_above", time_above, "min", symbols=["mid"])],
    keep=["t_above"],
)
print(res.q["t_above"].to("hr").round(1))
Keep only what you need
keep lists the observables in the result, all of them by default. The id of a PK observable keeps all of its parameters. Other observables are computed where a kept one needs them and then dropped, and an observable that no kept one needs is not computed at all:
res = simulator.run(MIDAZOLAM_SBML, doses, observables, keep=["pk"])
print(len(res.variables), res.variables[:3], "time" in res.ds.dims)
A result whose kept observables are all values per simulation has no time dimension. This saves a lot of memory in a large scan: 100,000 points with 481 time points and two timecourses take 0.8 GB, the same points with ten values per simulation 8 MB.
keep may also name variables of the model, e.g. keep=["pk.cmax", "[Cve_mid]"]. A kept variable is a timecourse under its own name and in the unit of the model, so one run gives both the values per simulation and the timecourses for a figure.
Errors
With on_error="flag", a point that fails gets NaN for every observable and the variable status marks it, see Parameter scans; res.nca gives it the flag NCAFlag.NO_DATA. A point fails exactly when its own simulation or its own observables fail. With on_error="raise" (the default), the run raises for the first failing point in the order of the scan.